AI-Driven Drug Discovery
Integrating machine learning and predictive architectures to accelerate lead identification, drug repurposing, and candidate prioritization.
From AI-driven virtual screening to multi-omics data integration, we deliver agile computational biology solutions that accelerate therapeutic discovery, optimize lead candidates, and drive publication-quality scientific outcomes.
End-to-end computational biology pipelines engineered to de-risk and accelerate therapeutic discovery.
Machine-learning virtual screening & predictive lead modeling
Molecular docking, cavity mapping & 100ns+ MD trajectories
NGS RNA-Seq, proteomic profiling & pathway networks
Binding free energy calculations & per-residue decomposition
Linux HPC workflows, Python/R automation & reproducible code
ProGen Advance Insilico Solutions LLP isn't just another bioinformatics service provider. Founded to bridge computational intelligence and molecular biophysics, we combine cutting-edge AI architectures with a pragmatic, reproducible scientific approach.
Our multidisciplinary team works side-by-side with clients and academic researchers to tackle complex drug discovery, genomic analysis, and molecular modeling challenges — turning biological complexity into opportunities for scientific breakthrough.
End-to-end in silico capabilities spanning therapeutic discovery, biophysical modeling, systems biology, and custom research pipelines.
Integrating machine learning and predictive architectures to accelerate lead identification, drug repurposing, and candidate prioritization.
Target structure modeling, high-throughput virtual screening, 100ns+ all-atom molecular dynamics, and MM/GBSA binding free energies.
Processing high-throughput genomics, transcriptomics, proteomics, and pathway enrichment datasets to identify robust biomarkers.
Developing automated bioinformatics pipelines, reproducible Linux-based computational workflows, and custom analytics scripts.
Real computational projects demonstrating the integration of AI, molecular modeling, and multi-omics analysis.
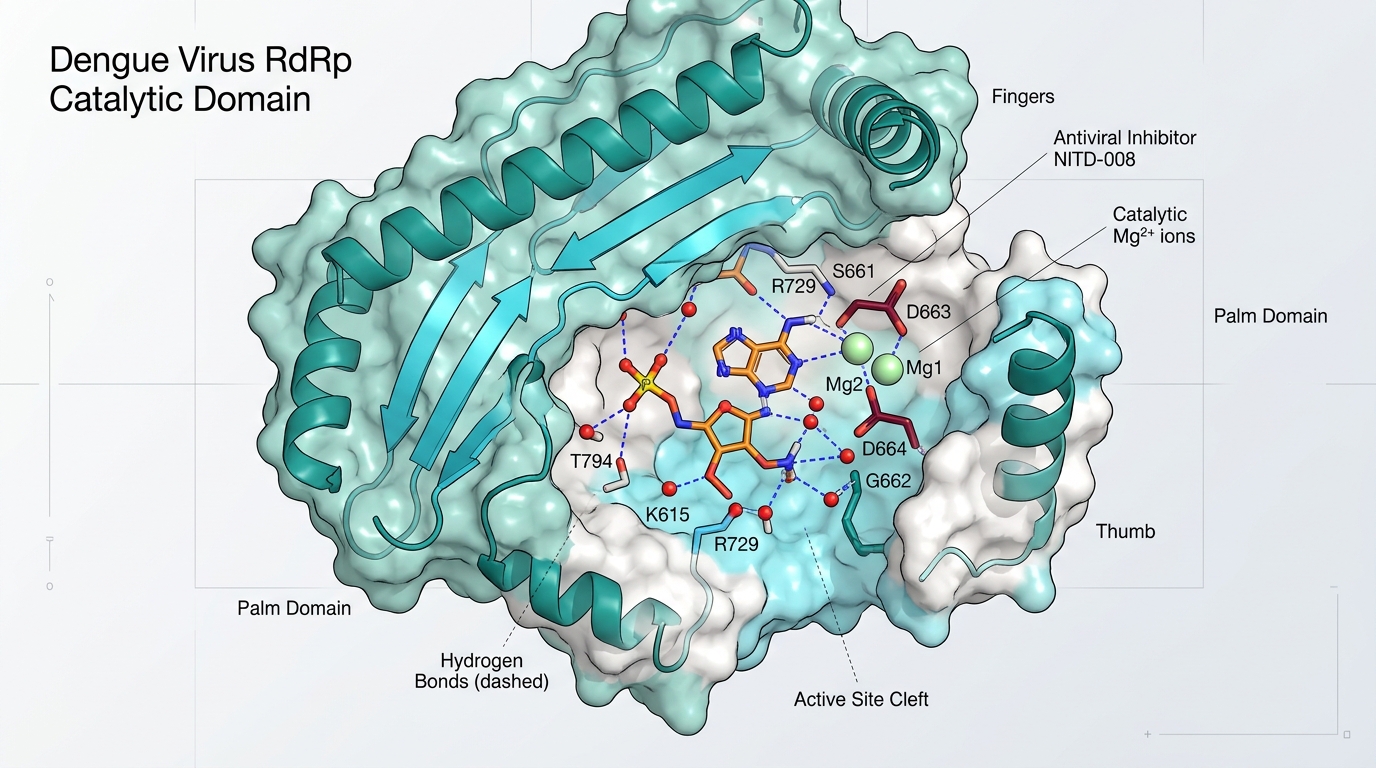
Dengue virus remains a global health threat with limited antiviral options. We characterized the conserved RNA-dependent RNA polymerase (RdRp) catalytic domain, screened diverse chemical libraries using RDKit similarity search, and evaluated binding energetics via 100ns molecular dynamics.
Interaction stability and binding free energies (MM/GBSA) proved substantially more reliable indicators of antiviral potency than docking scores alone.
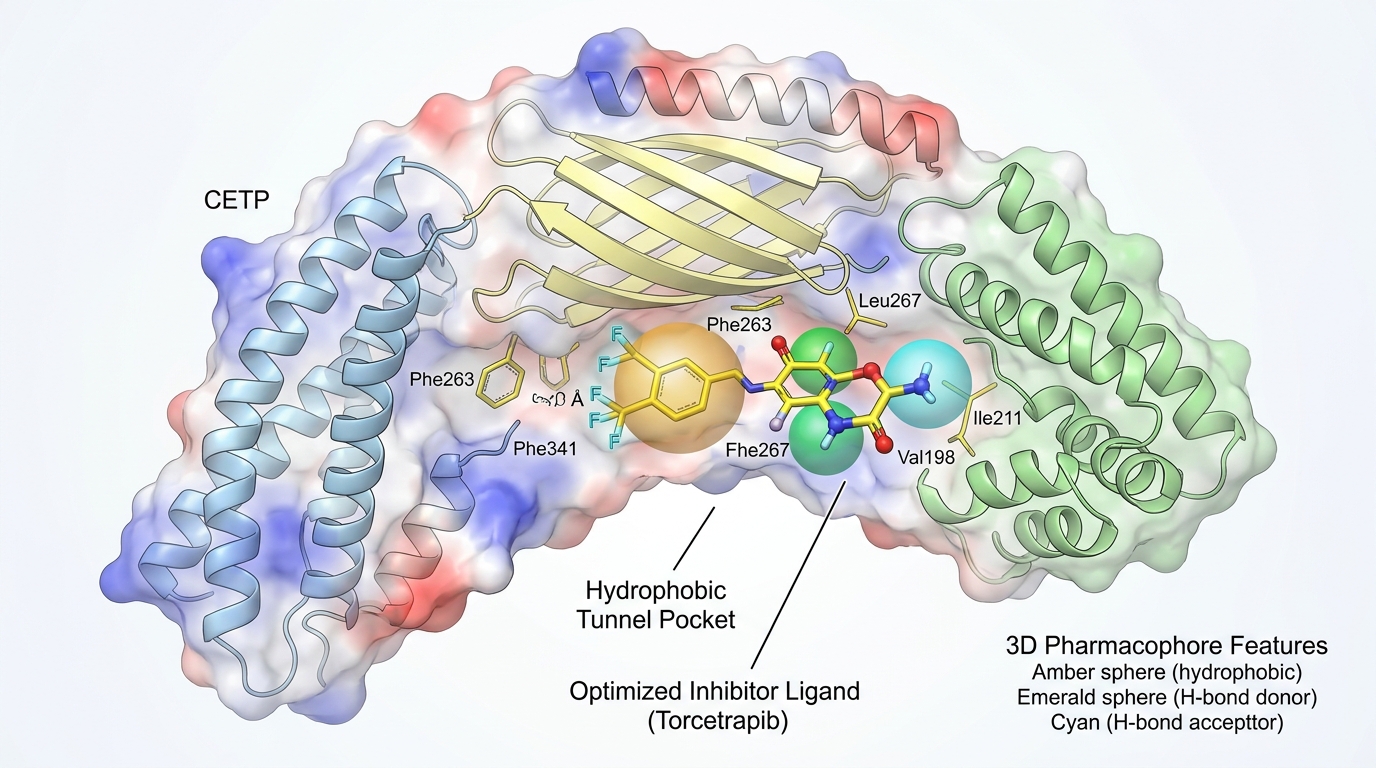
Cholesteryl Ester Transfer Protein (CETP) plays a central role in lipid metabolism. By combining pharmacophore modeling with a high-accuracy machine learning classifier (Score: 0.87), we evaluated sustained interactions within the CETP tunnel region.
Simulations mapped critical stabilizing residues within the hydrophobic tunnel, establishing a validated pipeline for structure-guided lead prioritization.
From biological question formulation to reproducible data preparation, AI modeling, and actionable outcomes.
Defining disease mechanisms, target selection, and scientific hypotheses.
Curation of PDB structures, NGS read sets, and curated chemical libraries.
Virtual screening, molecular dynamics, and predictive deep learning runs.
Translating algorithmic binding scores into biological pathway contexts.
Prioritized leads, biomarker signatures, and publication-ready dossiers.
Supporting researchers, innovators, and organizations across the life science and biomedical ecosystem.
MSc, PhD, and postdoctoral scholars receiving publication-grade analysis and structural modeling.
Target identification, high-throughput virtual screening, and structure-based lead optimization.
Custom automated bioinformatics pipelines, AI modeling, and multi-omics data integration.
Genomic variant analysis, biomarker discovery, and neoantigen immunogenicity prediction.
Computational research component execution and multi-laboratory consortium support.
Affordable, agile in silico screening and technical feasibility validation prior to wet-lab synthesis.